From Semaglutide to Oral Macrocycles: How Noncanonical Amino Acids Transform Peptide Medicines
Using seven approved peptide drugs as examples, this article explains how Aib, D-amino acids, N-methylation, fatty diacids, and other specialized residues work together to improve stability, conformation, target recognition, and pharmacokinetics.
Natural proteins primarily use 20 common amino acids, but drugs do not need to be constrained by this “alphabet.” For peptides, a natural sequence is often only the starting point for biological activity: it may be rapidly cleaved by proteases, have too short a circulating half-life, be overly conformationally flexible, or have difficulty crossing biological membranes because of high polarity. Medicinal chemists introduce unnatural amino acids not to make molecules look more complex, but to alter stereochemistry, backbone conformation, side-chain interactions, and metabolic pathways at specific positions.
Marketed peptide drugs provide the clearest evidence. From the Aib in Semaglutide and Tirzepatide, to the D-amino acids in Octreotide, backbone N-methylation in Cyclosporine A, and the oral macrocyclic peptide Enlicitide and the natural product Daptomycin, these molecules do not use the same chemical strategies, yet together they illustrate one point: whether a residue is valuable depends on where it is placed, what problem it solves, and whether it can act synergistically with the rest of the molecule’s design.
Why Are the 20 Natural Amino Acids Still Not Enough?
Proteases have evolved to recognize natural peptide backbones and common side chains; natural peptides also typically need to retain a certain degree of flexibility to perform physiological functions. Once they enter drug development, these characteristics can become weaknesses. A D-configuration can change the three-dimensional geometry seen by enzymes and receptors; α,α-disubstituted residues can restrict the conformations accessible to the backbone; N-methylation can remove a hydrogen-bond donor and alter the cis–trans equilibrium of a peptide bond; larger aromatic side chains, charged side chains, or special residues that can be used for cyclization can access binding space that is difficult for natural amino acids to cover.
These changes do not automatically produce a “better drug.” A more stable conformation may also lead to loss of activity or solubility; increased hydrophobicity may enhance albumin binding, but may also cause aggregation or nonspecific binding; reducing hydrogen-bond donors can sometimes favor membrane permeability, but in other cases can disrupt receptor recognition. The significance of unnatural amino acids therefore lies not in their number, but in enabling more precise tuning of molecular properties.
Aib: The Change Brought by One Methyl Group
Aib, or α-aminoisobutyric acid, appears to differ from Alanine only by an additional methyl group on the α-carbon. This change makes the α-carbon an α,α-disubstituted center bearing two identical methyl groups, so it is no longer chiral, while also markedly changing the steric environment and conformational preferences around the backbone. Whether a protease can approach a cleavage site, and which local conformations the peptide chain tends to adopt, may both change as a result.
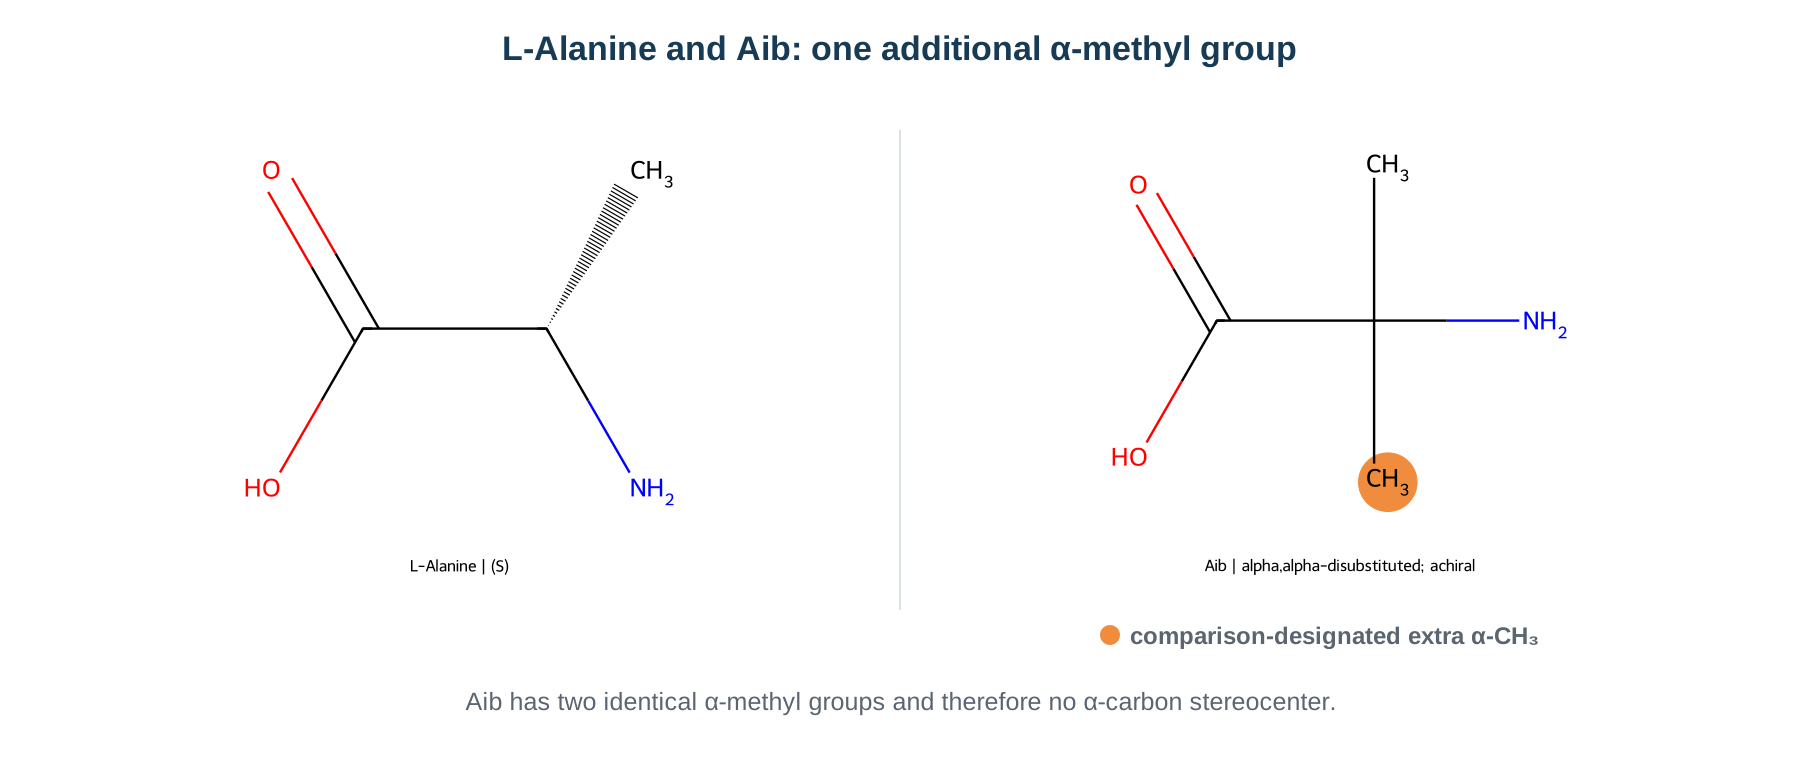
Figure 1. Validated two-dimensional structures of L-Alanine and Aib. The two α-methyl groups in Aib are identical, so its α-carbon is not a chiral center; the orange color only indicates the structural comparison relative to Alanine. Structure sources: PubChem CID 5950 and 6119, drawn deterministically using RDKit.
Semaglutide is one of the best-known examples of this design. Ala at position 8 of natural GLP-1 is located in a region readily cleaved by DPP-4; replacing it with Aib can enhance stability against DPP-4 degradation. However, once-weekly dosing cannot be attributed solely to Aib. Semaglutide also incorporates a C18 fatty diacid on the Lys26 side chain through two OEG units and a glutamic acid-derived linker unit, enhancing reversible serum albumin binding; Lys at position 34 is replaced with Arg, allowing the fatty diacid to be selectively conjugated at Lys26. FDA materials identify albumin binding as a major long-acting mechanism and report an elimination half-life of approximately one week. In other words, Aib8 primarily addresses a protease-labile site, while the Lys26–linker–C18 diacid primarily addresses albumin binding and in vivo retention; the two effects are different but complementary.
Liraglutide provides a meaningful comparison. It is also lipidated at Lys26, but uses a glutamic acid spacer to attach a C16 fatty acid, with a plasma half-life of approximately 13 hours, making it suitable for once-daily dosing. The C18 diacid and longer hydrophilic linker of Semaglutide strengthen albumin binding, and Aib8 also increases stability against DPP-4. The difference between daily and weekly dosing therefore cannot be simplified as “C16 versus C18”; resistance to enzymatic degradation, linker structure, albumin binding, self-association, and overall clearance all jointly determine dosing frequency.
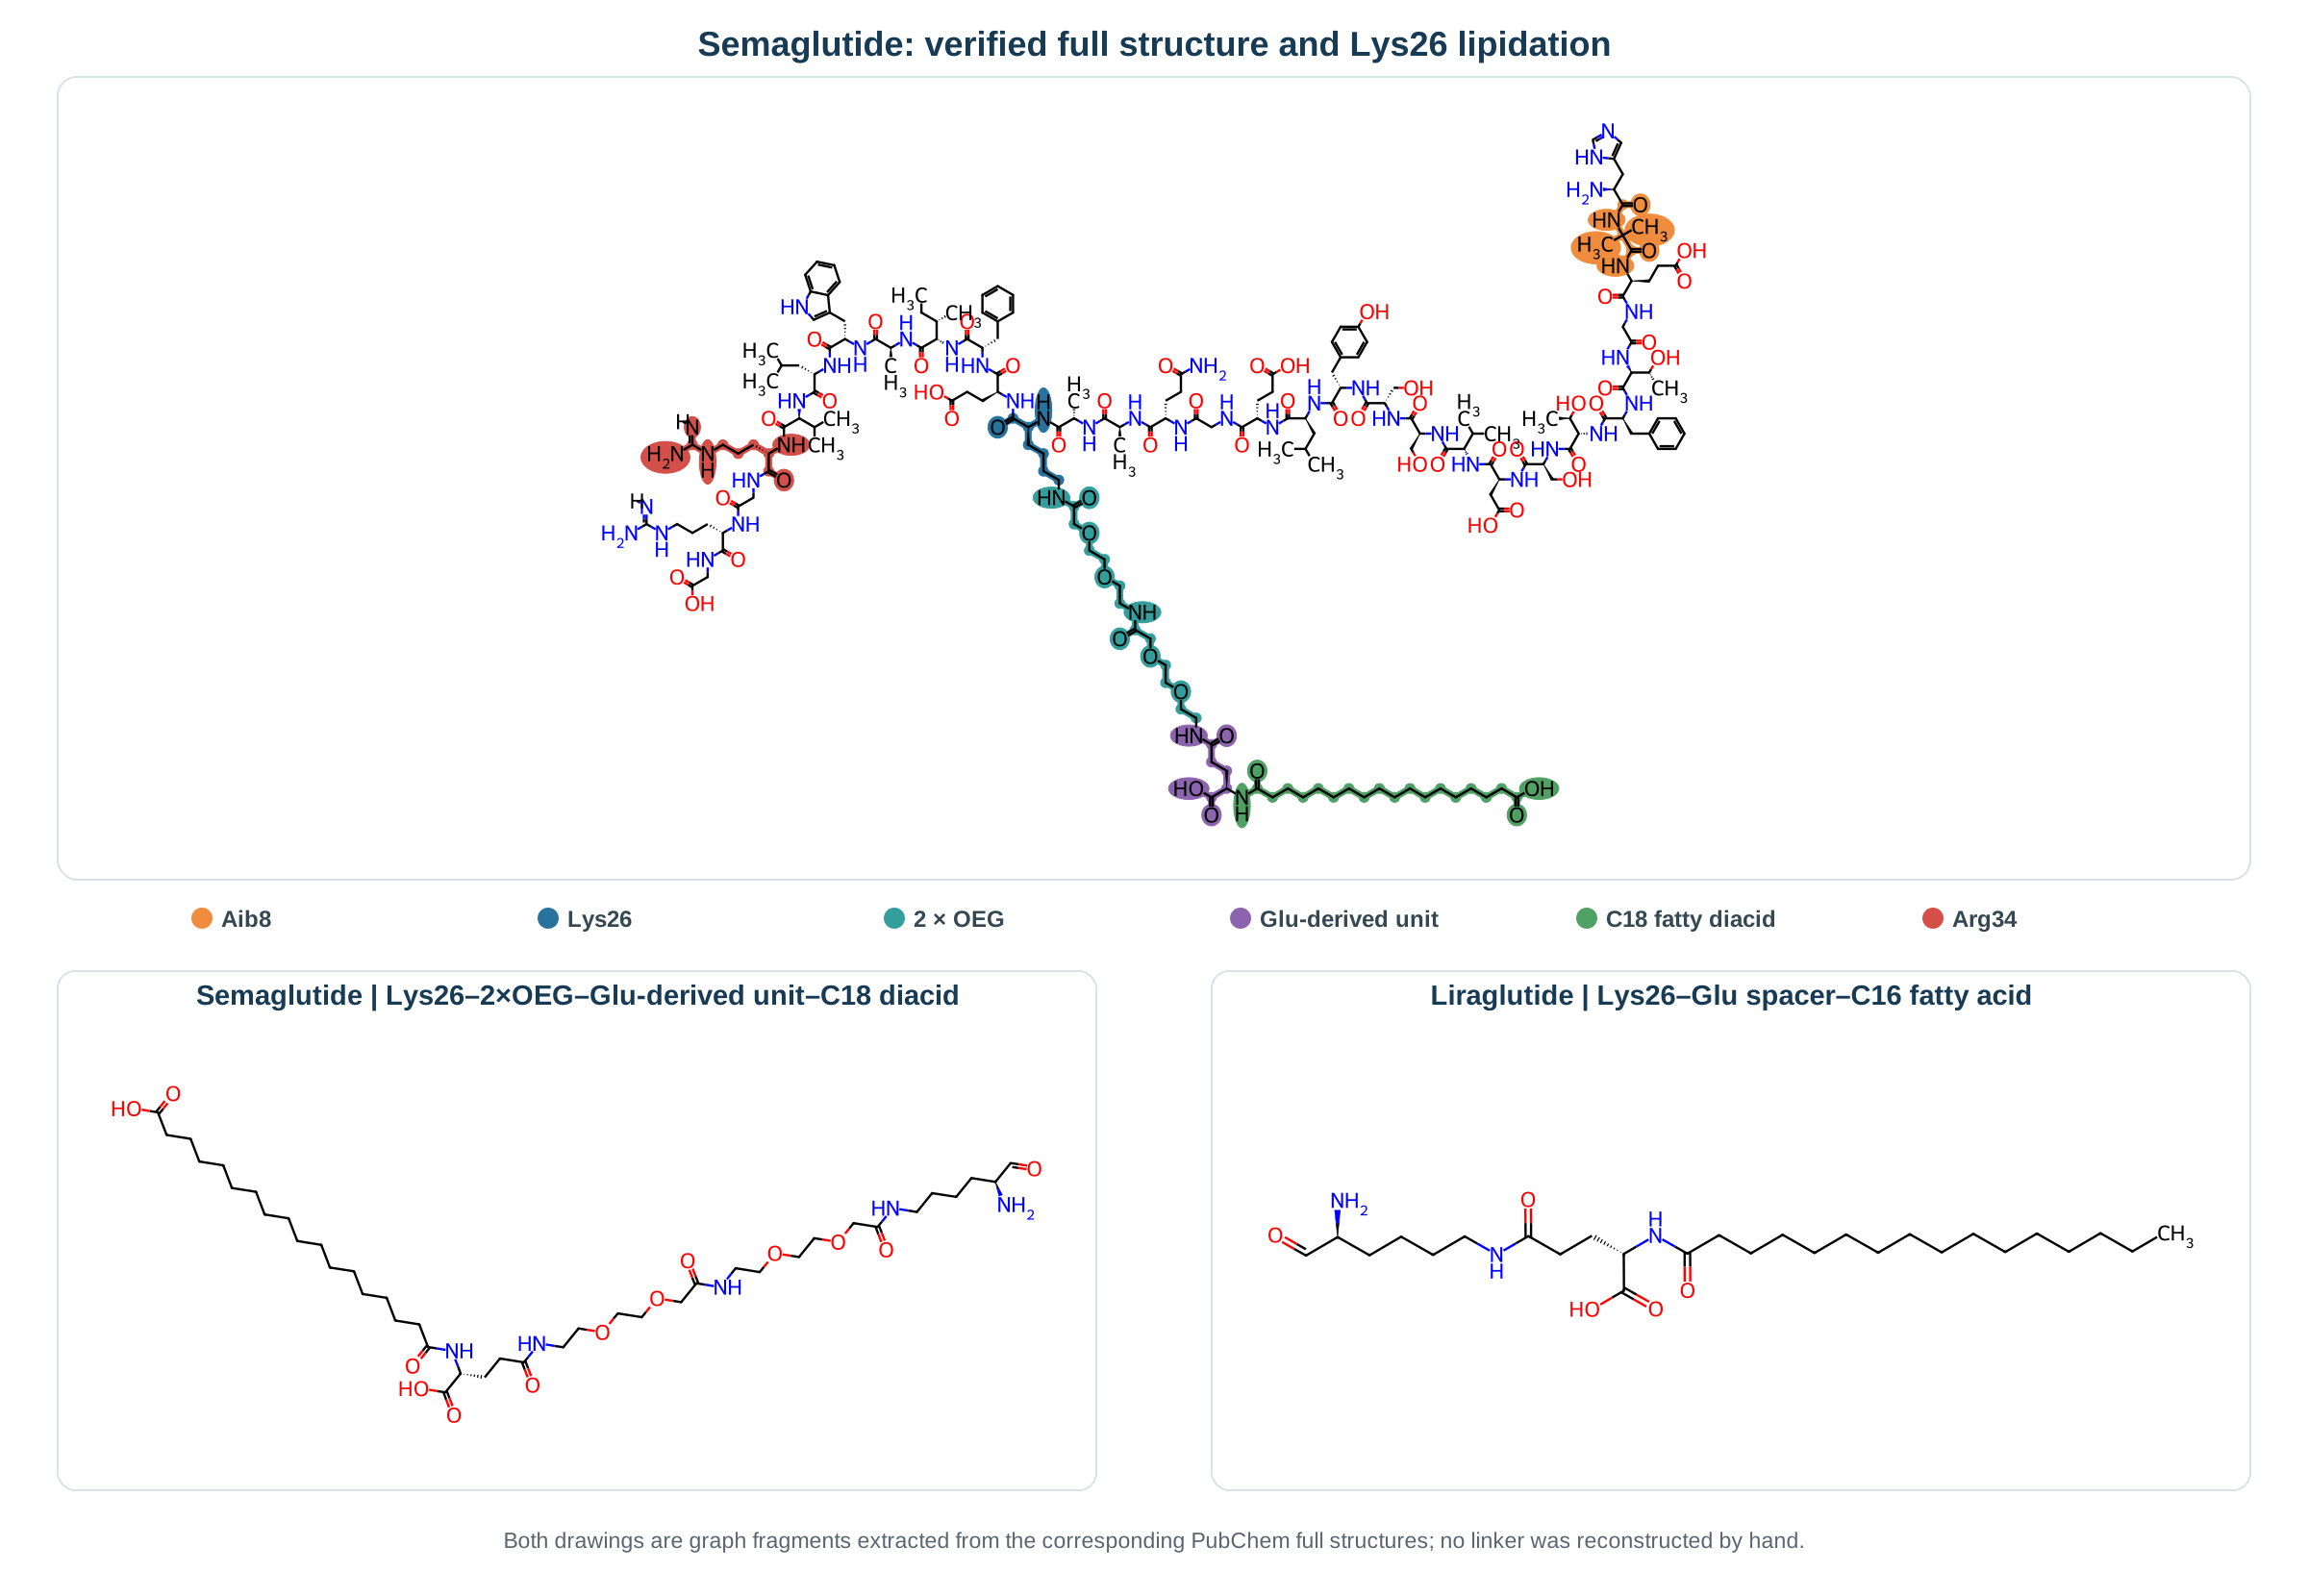
Figure 2. Complete structure of Semaglutide and the Lys26 modification region, compared with the local lipidation structure of Liraglutide. The complete molecules are from PubChem CID 56843331 and 16134956, respectively; the local structures were extracted directly from the corresponding full-molecule diagrams without manual splicing. Aib8, Lys26, and Arg34 in Semaglutide were localized by mapping to the known GLP-1 sequence, and the OEG units, glutamic acid-derived unit, and C18 diacid were each uniquely matched as substructures within the full molecule.
Tirzepatide applies a similar strategy to a long-acting GIP/GLP-1 dual receptor agonist. The FDA prescribing information shows that it contains two Aib residues at positions 2 and 13, and incorporates a C20 fatty diacid at Lys20 through a linker. Aib, lipidation, and the adjusted peptide sequence together support protease stability, albumin binding, and an approximately 5-day half-life, ultimately enabling once-weekly dosing. Here as well, no single modification should be presented as the entire answer: dual-receptor pharmacology comes from the whole sequence, while long-acting properties arise from a combination of multiple structural designs.
D-Amino Acids: Changing Stereochemical Configuration, Not Just “Preventing Degradation”
Octreotide is a classic marketed drug containing D-amino acids. It is a disulfide-bridged octapeptide whose sequence contains D-Phe and D-Trp. Compared with natural somatostatin, it is shorter and more stable, while retaining pharmacological activity at somatostatin receptors. Structural studies show that the D-Trp–Lys region can form a specific β-turn, and the spatial relationship among D-Phe, D-Trp, and Lys is also an important component of receptor recognition.
This shows that the role of D-amino acids should not be simply summarized as “all D-residues are more stable.” Inverting the configuration of an α-carbon simultaneously changes the trajectory of the backbone and the orientation of the side chain; if the position is appropriate, it may evade protease recognition while preserving a favorable conformation, but if the position is inappropriate, it may directly impair affinity. The success of Octreotide comes from stereochemical coordination within a specific sequence, not from a rule that can be copied arbitrarily.
N-Methyl Amino Acids: Lessons from Cyclosporine A
Cyclosporine A is a cyclic natural product composed of 11 residues and contains D-Ala and seven amide N-methylation sites. N-methylation converts backbone N–H into N–CH3, reducing hydrogen-bond donors and altering local steric hindrance, peptide-bond cis–trans isomerization, and overall conformational energy. For a macrocyclic molecule, these changes rearrange which polar groups are exposed to solvent and which can be temporarily “hidden” by intramolecular hydrogen bonds.
Cyclosporine A is often described as having “chameleonic” conformational properties: in environments of different polarity, it can reorganize its conformation and hydrogen-bonding network, thereby changing apparent polar exposure. This provides an important example of how macrocyclic peptides can cross membrane barriers, but it does not justify the conclusion that “N-methylation necessarily improves membrane permeability.” When placed at inappropriate positions, N-methylation may disrupt key hydrogen bonds, alter the binding conformation, or even reduce solubility. What truly matters is the coordination among the N-methylation pattern, D-residues, ring size, and the entire conformational ensemble.
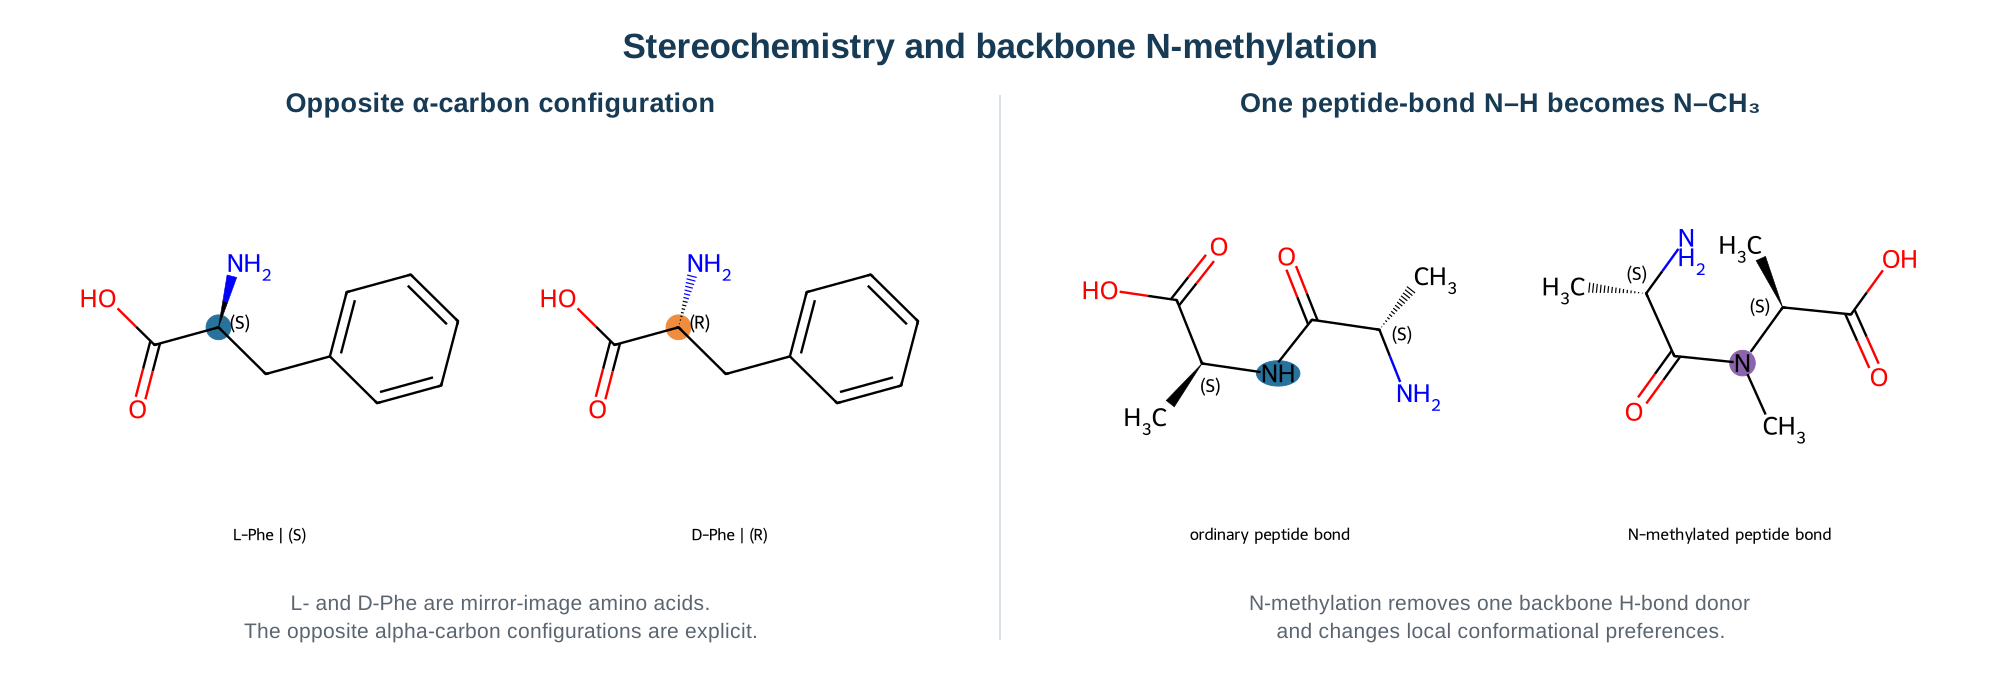
Figure 3. Left: Mirror-image configurations of PubChem L-Phe (CID 6140) and D-Phe (CID 71567). Right: A conventional dipeptide model and an N-methylated dipeptide model, highlighting replacement of one backbone hydrogen-bond donor by a methyl group. Structures were drawn using RDKit with explicit stereochemistry retained.
From Special Side Chains to the Oral Macrocyclic Peptide Enlicitide
Enlicitide (development code MK-0616) takes the role of unnatural amino acids to a more complex level. In early PCSK9 macrocyclic peptides, 5-fluorotryptophan expanded aromatic side-chain chemistry, and α-methylproline changed local conformation; subsequent studies then used structure-guided D-amino acid substitution, backbone N-methylation, and multiple cyclizations to progressively address protease stability and conformational preorganization. These modifications did not proceed in a straight line, nor did each one bring only benefits: the five-part Enlicitide series already on this website documents how improvements in stability continued to expose issues in solubility, clearance, formulation, and manufacturing.
The significance of the final molecule lies not in “using many unnatural amino acids,” but in different structural elements separately contributing to target binding, conformational restriction, stability, and optimization of overall drug properties. A 2026 development paper also shows that, to avoid problems such as oxidation, low solubility, and process safety, the team systematically reconstructed late-stage scaffolds and synthetic building blocks rather than simply retaining the early hit. To learn more about the process from a high-affinity tricyclic lead to an oral candidate, continue reading the oral PoC stage and formation of the final candidate.
The regulatory status also needs to be described according to the latest information. As of October 2026, Enlicitide is no longer an investigational-stage candidate. The FDA approved an oral tablet under the trade name LIPFENDRA in July 2026; the label uses the generic name enlicitide, and its drug substance is enlicitide decanoate, indicated as an adjunct to diet and exercise to reduce LDL-C in adult patients with hypercholesterolemia. It is a once-daily oral PCSK9 inhibitor. The molecule’s unnatural chemistry addresses part of the challenges in binding, stability, and developability, while the approved product still also relies on oral formulation design; therefore, “containing unnatural amino acids” and “being orally available” cannot be treated as simply equivalent. For differences between the two approved oral cyclic peptides, see “The Different Paths of Icotrokinra and Enlicitide”.
Natural Products Have Long Used an “Expanded Alphabet”
Daptomycin is a cyclic lipopeptide antibiotic produced by microorganisms, composed of 13 residues and a decanoyl fatty acyl chain. The FDA chemical name and PubChem structure show that it contains D-Asn, D-Ala, and D-Ser, as well as Ornithine, Kynurenine, and 3-methylglutamic acid; the 10 C-terminal residues form an ester-linked macrocycle. These special residues do not merely make the molecule more difficult for proteases to process; together with the fatty acyl chain, cyclization mode, and acidic groups, they shape the molecule’s conformation, calcium-ion response, and membrane interactions.
The antibacterial mechanism of Daptomycin remains under active investigation, but calcium-dependent membrane binding and effects on phosphatidylglycerol-rich Gram-positive bacterial membranes are central elements in the public evidence. It is inaccurate to designate any one special residue alone as the entire source of pharmacological activity; the lesson from natural products is precisely that unnatural residues, lipidation, and macrocyclic topology can jointly produce function within an integrated structure.
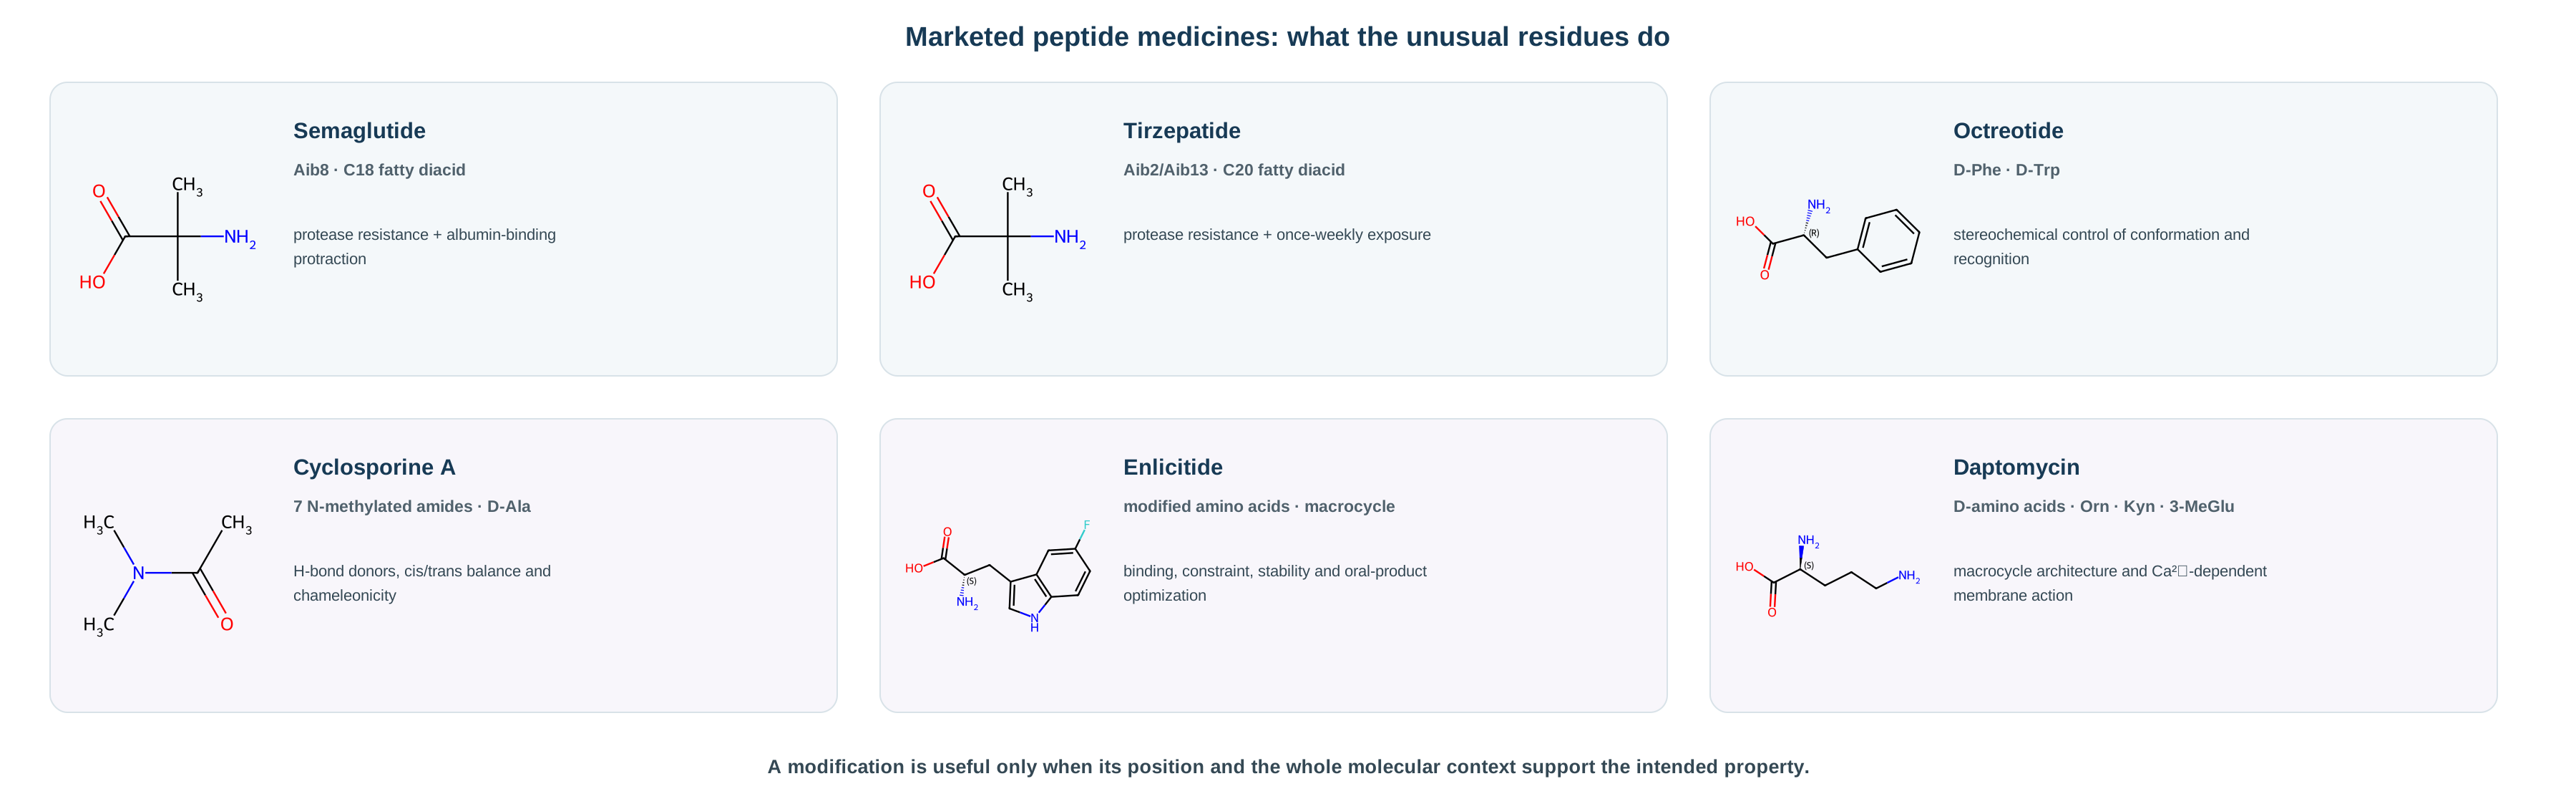
Figure 4. Representative non-natural chemistries in six marketed peptide drugs. The small structural fragments are derived from verified Aib, D-Phe, N-methyl amide, 5F-Trp, and Orn structures and are used only to illustrate residue classes; they do not replace the complete structures of the respective drugs.
From Single-Residue Replacement to Overall Molecular Design
These marketed drugs do not provide a universal “best list of non-natural amino acids.” Semaglutide and Tirzepatide show that a protected cleavage site must work in concert with lipidation, a linker, and the entire receptor-agonist sequence; Octreotide shows that stereochemical configuration itself can participate in receptor recognition; Cyclosporine A illustrates the relationship between N-methylation patterns and dynamic conformations; and Enlicitide and Daptomycin remind us that the function of complex macrocycles arises from the combined effects of residues, cyclization, and overall molecular properties.
This preserves the central point of the earlier article: a noncanonical amino acid is not an isolated “stability plug-in,” but a design variable whose value must be judged in the context of the whole medicine.
Therefore, the value of non-natural amino acids lies not in how many types are used, but in whether the right positions and structures can be used to solve clearly defined problems. mRNA Display can directly bring this type of chemistry into ultra-large-scale screening, expanding the discovery space of cyclic peptides; chemical synthesis can convert screening leads into real samples; and computational and structure-based design helps propose more focused substitution strategies. Ultimately, however, whether a design is valid is still determined by synthesis, structural characterization, and experiments on binding, function, stability, and pharmacokinetics.
References
- Lau J, Bloch P, Schäffer L, et al. Discovery of the Once-Weekly Glucagon-Like Peptide-1 (GLP-1) Analogue Semaglutide. Journal of Medicinal Chemistry. 2015;58:7370–7380. https://doi.org/10.1021/acs.jmedchem.5b00726
- U.S. FDA. WEGOVY (semaglutide) Prescribing Information. FDA label
- U.S. FDA. VICTOZA (liraglutide) Prescribing Information. FDA label
- U.S. FDA. ZEPBOUND (tirzepatide) Prescribing Information. FDA label
- U.S. FDA. SANDOSTATIN (octreotide acetate) Prescribing Information. FDA label
- Ono S, et al. Cyclosporin A: Conformational Complexity and Chameleonicity. Journal of Chemical Information and Modeling. 2021. PubMed PMID 34672629
- Tucker TJ, Embrey MW, Alleyne C, et al. A Series of Novel, Highly Potent, and Orally Bioavailable Next-Generation Tricyclic Peptide PCSK9 Inhibitors. Journal of Medicinal Chemistry. 2021;64:16770–16800. https://doi.org/10.1021/acs.jmedchem.1c01599
- Josien H, Nair AG, Ding F-X, et al. Discovery Process of Enlicitide, a Highly Engineered Macrocyclic Peptide Therapeutic, through Issue-Driven Fragment-Based Synthetic Assembly and SAR. Journal of Medicinal Chemistry. 2026;69:13473–13491. https://doi.org/10.1021/acs.jmedchem.6c00463
- U.S. FDA. LIPFENDRA (enlicitide) Prescribing Information. FDA label
- U.S. FDA. CUBICIN (daptomycin) Prescribing Information. FDA label
Need to evaluate a synthesis and validation plan for a noncanonical peptide?
If your project involves Aib, D-amino acids, N-methyl amino acids, lipidation, or complex macrocycles, we can support structure review, custom synthesis, purification, and quality analysis.